CompPhy
CompPhy: a web-based collaborative platform for comparing phylogenies
CompPhy is a web platform dedicated to the collaborative handling of phylogenetic trees. Users can freely manage collections of trees and communicate on a common project. By collaborative, we mean that several users connected to the same project can manipulation at the same time trees from shared collections. The platform is flexible enough to allow each user to focus on some trees or to follow trees manipulated by another user. The platform offers various functionalities covering tree edition, tree comparison, supertree inference, data management.
Why CompPhy?
Online collaborative and real-time tools are needed in various domains for projects involving remote partners. In the last twenty years, web technologies have enabled the development of such tools to jointly edit office documents. However, only few such tools exist in scientific domains. Particularly in phylogentics, prior to the introduction of the CompPhy website, no collaborative tool was available to handle phylogenies.
Key features
- Collaborative editing collection of trees: shared tree visualization, (a)synchronous manipulation of trees, data exchange/storage, …
- Tree edition and taxa annotation: coloration and fonts, shape of the tree, subtree collapsing, …
- Tree comparison: side-by-side display, swapping leaves to ease comparison, MAST, Robinson & Foulds distance
- Supertree inference: MRP, Physic_IST, Astral
- Data management: timeline, associated documents, message forum, backups, …
Access the tool
Use CompPhy here: http://old.atgc-montpellier.fr/compphy/
User guide
Consult the user guide to discover all functionalities.
Scientific reference
« CompPhy: a web-based Collaborative Platform for Comparing Phylogenies », N. Fiorini, V. Lefort, F. Chevenet, V. Berry, A.-M. Arigon Chifolleau, BMC Evol Biol. 2014 Dec 14; 14(1):253.
Other tools
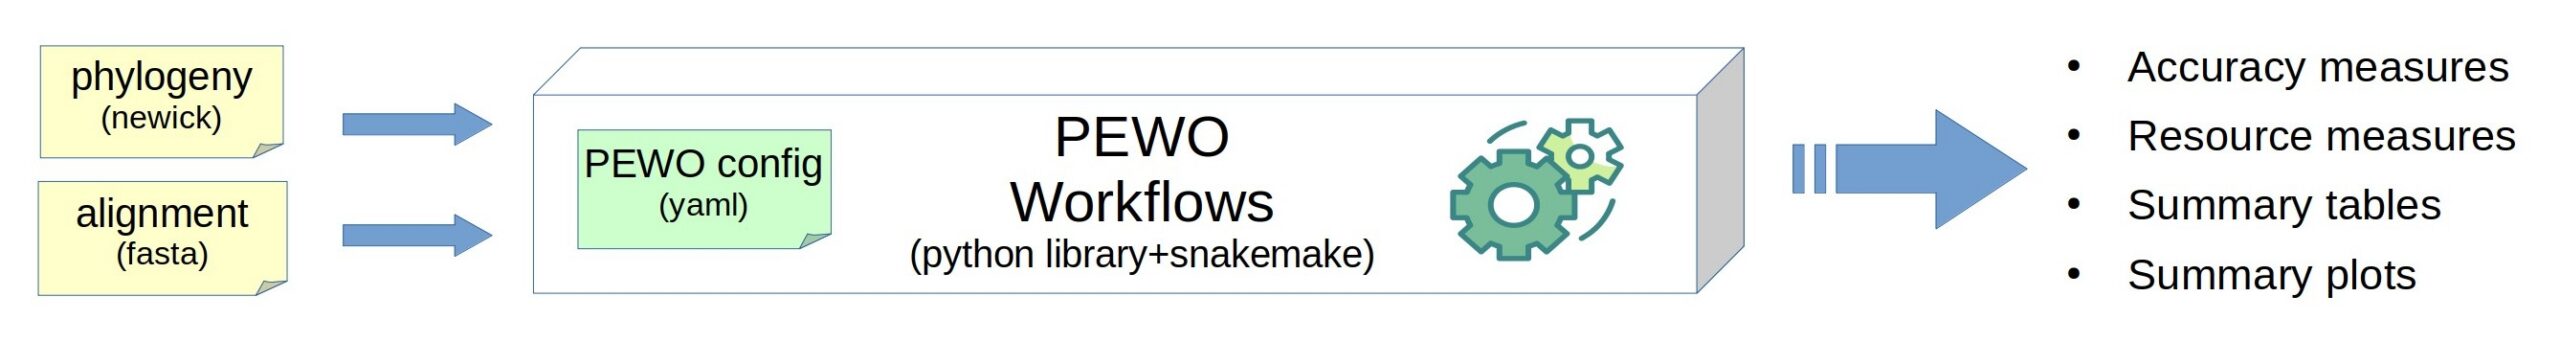
PEWO: a collection of workflows…
Introduction and context In the Bioinformatics team of the LIRMM (CNRS & Univ. Montpellier), we develop a series of tools for metagenomics / metabarcoding analysis. Our tools exploit phylo-k-mers (which are k-mers combined with phylogenetic information) computed for an input set of reference sequences and their phylogeny. The phylo-k-mers are computed and indexed with IPK,…
DExTER
Overview DExTER (Domain Exploration To Explain gene Regulation) is a bioinformatics tool designed to automatically identify genomic regions whose nucleotide composition correlates with gene expression levels. Unlike traditional approaches focusing on short transcription factor binding sites (6-12 bp), DExTER detects Long Regulatory Elements (LREs) that can span tens to hundreds of nucleotides. This makes it…
LoRDEC: hybrid correction of long…
Overview In a nutshell, LoRDEC is a program for error correcting long sequencing reads using short reads. It implements a hybrid correction approach. It uses little memory and is very efficient. Most importantly it scales up to process very large data sets. It can be applied to long reads obtained with either Pacific Biosciences SMRT…