SMS: Smart Model Selection in PhyML
SMS (Smart Model Selection) automatically selects the best-fitting evolutionary model for an alignment of DNA or protein sequences. It compares substitution models using the AIC or BIC statistical criteria to support phylogenetic tree reconstruction with PhyML.
SMS: Smart Model Selection in PhyML online execution
Other tools
CompPhy
CompPhy: a web-based collaborative platform for comparing phylogenies CompPhy is a web platform dedicated to the collaborative handling of phylogenetic trees. Users can freely manage collections of trees and communicate on a common project. By collaborative, we mean that several users connected to the same project can manipulation at the same time trees from shared…
FastME
FastME is a software package for the fast and accurate inference of phylogenetic trees from distance matrices. It implements algorithms based on the Balanced Minimum Evolution (BME) principle, a distance-based criterion closely related to the Neighbor Joining (NJ) method. The goal of the BME framework is to identify the phylogenetic tree that minimizes the total…
RSCU_RS: Measuring the bias in…
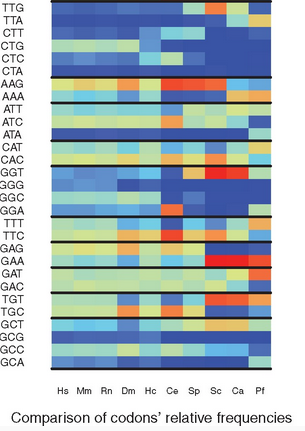
Overview Overview: In the protein coding sequences of a species, the 61 possible codons of the genetic code are not equally distributed. This observation is referred to as the Codon Usage Bias (CUB) of a species. Several measures have been proposed to quantify the CUB using the frequencies of codons in all RNA coding sequences…