RSCU_RS: Measuring the bias in codon usage from ribosomal activity
Overview
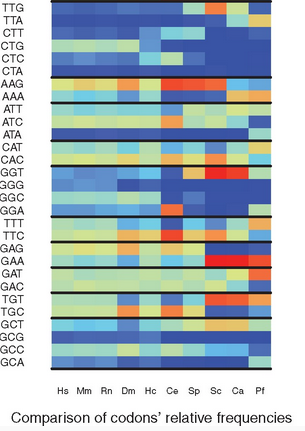
Overview: In the protein coding sequences of a species, the 61 possible codons of the genetic code are not equally distributed. This observation is referred to as the Codon Usage Bias (CUB) of a species. Several measures have been proposed to quantify the CUB using the frequencies of codons in all RNA coding sequences (or at least a representative subset of these). This yields a static analysis since sequences change slowly over time. But in living cells the translation varies over time, since the ribosome, the molecular machine that performs translation, has a role in the selection of which RNA sequences are indeed translated. The precise location of ribosomes translating RNA can be monitor by Ribo-seq (aka ribosome profiling), a high throughput sequencing assay that captures the portions of RNA located inside the ribosomes. Thus, the sequencing reads produced by Ribo-seq in a given condition give us access to which codons ribosomes are translating.
We proposed to measure the codon usage bias in a transcriptome wide manner using Ribo-seq sequencing data. This delivers a dynamic and precise estimation of Codon Usage Bias, since it integrates the location of ribosomes during translation. The codon usage bias can be computed classicaly for a given species, for a subset of genes, but also for any given condition for which Ribo-seq data is available. The CUB can thus be compared across species, across subsets of genes, or across conditions.
For this, we develop a stand-alone software, called RSCU_RS, and report experiments in estimating and comparing CUB across species in a journal article.
Reference
Journal article
Ribo-seq enlightens Codon Usage Bias
D. Paulet, A. David, E. Rivals
DNA Research, dsw062. https://doi.org/10.1093/dnares/dsw062, 2017.
Documentation and ressources
A documentation about the software RSCU_RS is available at
https://www.lirmm.fr/~rivals/rscu/
Funding
Fondation pour la Recherche Médicale, grant DBI20131228574.
ANR grant (ANR-11-BINF-0002), Institut de Biologie Computationnelle (IBC).
Other tools
WAVES
Warning : This application is no longer maintained and out of date. If you’re interested in, just have a look to MYST Summary WAVES is a web application dedicated to bioinformatic tool integration. It provides an efficient way to implement a service for any bioinformatic software. Such services are automatically made available in three ways:…
LoRDEC: hybrid correction of long…
Overview In a nutshell, LoRDEC is a program for error correcting long sequencing reads using short reads. It implements a hybrid correction approach. It uses little memory and is very efficient. Most importantly it scales up to process very large data sets. It can be applied to long reads obtained with either Pacific Biosciences SMRT…
PhyML
Overview: new algorithms, methods and utilities PhyML is a software package that uses modern statistical approaches to build phylogenetic trees from the analysis of alignments of nucleotide or amino acid sequences. The main tool in this package builds phylogenies under the maximum likelihood criterion. It implements a large number of substitution models coupled to efficient…